Why Your Business Needs a Custom Method Development Laboratory
Why a Custom Method Development Laboratory Matters
A custom method development laboratory builds and proves an analytical procedure when a standard method cannot reliably answer your question. This is essential when you need lower detection limits, must test a difficult or proprietary matrix, need to measure a new compound or trace impurity, or require data that can stand up to quality and regulatory review.
A strong project starts by defining the required result: the analyte, sample type, reporting range, detection limit, accuracy, turnaround needs, and validation level. The laboratory can then select the right sample preparation, separation, and detection approach – such as ICP-MS, LC-MS/MS, GC-MS, or chromatography – and test whether the method remains reliable across normal operating changes.
For aerospace, pharmaceutical, and environmental teams, this work reduces a common risk: receiving data from a method that was never designed for the real sample. Off-the-shelf methods can be useful starting points, but they may not address matrix interference, ultra-low concentrations, extraction losses, or unique regulatory requirements.
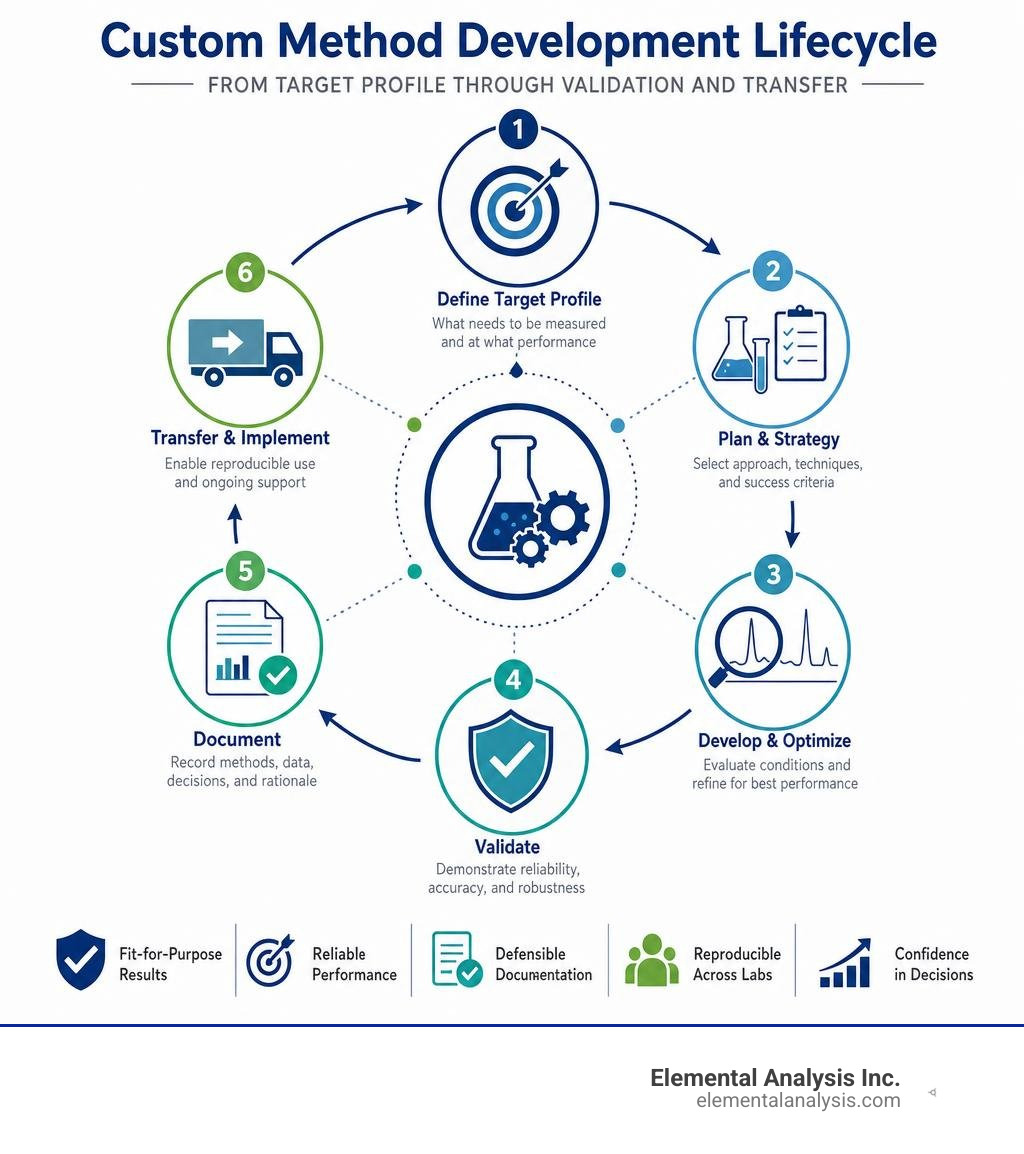
Custom development also creates a defensible record of how the method was designed, optimized, validated, and transferred. Clear, contemporaneous notes matter because small details – reagent lots, sample hold times, instrument settings, and rejected conditions – can determine whether another analyst or laboratory can reproduce the result.
Custom method development laboratory terms made easy:
- specialized analytical chemistry services
- troubleshooting analytical method problems
- accredited testing laboratory services
What Is Custom Method Development and When Is It Essential?
Every analytical testing workflow relies on a fundamental question: does the chosen test accurately quantify what is in the sample without interference from everything else? When standard, published protocols fail to meet this standard, custom method development bridges the gap.
Custom method development is the deliberate, experimental process of designing, refining, and testing a bespoke protocol to identify and quantify specific chemical or elemental targets. Rather than forcing an ill-fitting standard protocol onto a unique sample, custom development tailors every single variable—digestion chemistry, sample extraction, separation gradients, column chemistry, and detector tuning—to your target analytes and physical matrix.
This process establishes a fit-for-purpose testing protocol governed by an Analytical Target Profile (ATP). Defined formally under international frameworks like Master Lab Method Development: Step-by-Step Framework 2026 | Verbex, an ATP details the required measurement uncertainty, target resolution, dynamic range, and acceptable limits of quantitation before an analyst even powers on an instrument. If your team is evaluating whether standard protocols are enough, reviewing A Beginner Guide To Industry Standard Methods helps clarify when routine compendial testing ends and custom chemistry begins.
Standard Compendial Methods vs. Tailored Analytical Procedures
Standard compendial methods—such as those published by USP, ASTM, EPA, or ISO—serve as valuable benchmarks across industries. They provide harmonized baselines for routine materials. However, their scope is intentionally broad, which creates severe operational limits when applied to non-standard formulations:
- Scope Constraints: Compendial procedures are validated only for specific matrices defined in the monograph. Using an ASTM water testing method on an abrasive slurry or a complex polymer emulsion invalidates the underlying validation assumptions.
- Extraction Efficiency Deficits: Standard digestion or extraction protocols often yield poor recovery when applied to novel composite materials, unvalidated biological fluids, or specialized nanomaterials.
- Detection Limit Ceilings: Standard procedures frequently target regulatory pass/fail thresholds established decades ago, leaving them unable to reach parts-per-billion (ppb) or parts-per-trillion (ppt) levels demanded by modern manufacturing quality standards.
- Matrix Interference Blind Spots: Compendial methods rarely account for proprietary excipients, specialized coating polymers, or unusual elemental ratios that suppress analytical signals or create false positives.
Tailored analytical procedures eliminate these blind spots by designing sample preparation and instrumental detection around your material’s exact physical and chemical behavior.
Critical Triggers Requiring Custom Method Establishment
Knowing when to transition from routine testing to custom development prevents costly project delays, failed validation audits, and skewed datasets. The most frequent triggers include:
- Novel Chemical Entities (NCEs) and Modalities: When synthesizing new active ingredients, advanced nanomaterials, or novel bioconjugates, nonproprietary methods simply do not exist.
- Proprietary and Difficult Matrices: Highly viscous oils, high-salt brines, complex biofluids, and advanced engineering ceramics resist standard sample breakdown techniques.
- Trace Impurity Demands: Modern semiconductor, aerospace, and medical device standards require rigorous monitoring of ultratrace contaminants. Learning How Trace Impurities Analysis Keeps Modern Technology Safe And Pure underscores why standard parts-per-million screening is no longer sufficient.
- Miniaturized Sample Availability: When research constraints leave you with microliters of precious fluid or microgram material shavings, standard high-volume digestion methods must be redesigned for micro-scale extraction.
- Regulatory Escalation: Advancing a compound from early exploratory research into regulated preclinical or clinical trials demands tighter analytical boundaries and formal validation.
The Core Workflow: How a Custom Method Development Laboratory Builds Validated Assays
Building a validated method is an iterative, structured scientific journey. Complex assays often take weeks or months of systematic development, moving intentionally through distinct phases to eliminate guesswork.
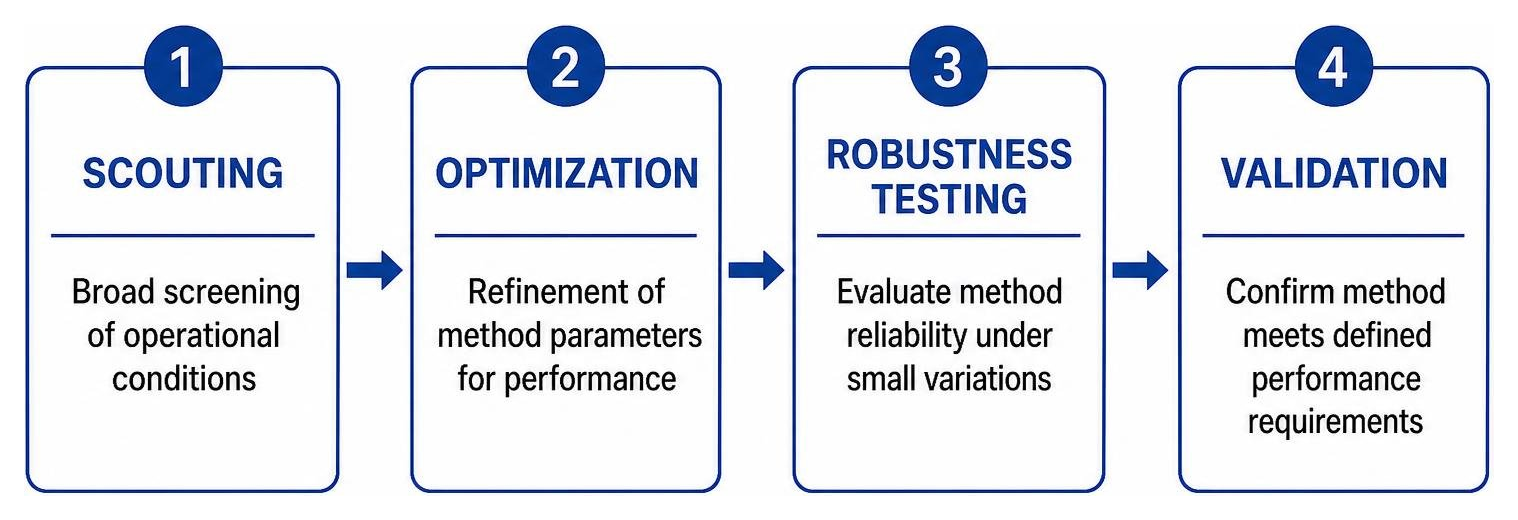
The table below outlines the core phases of development, tracking how analytical boundaries are tested from initial scoping to formal validation:
| Development Phase | Primary Objective | Key Experimental Parameters | Primary Deliverable / Milestone |
|---|---|---|---|
| Phase 1: Scouting | Broad screening of operational conditions | Stationary phases, solvents, pH, digestion acids, ionization modes | Identification of viable chemistry and retention mechanisms |
| Phase 2: Optimization | Maximizing resolution, recovery, and sensitivity | Gradient slopes, temperatures, extraction techniques, collision energies | Defined assay conditions and repeatable recovery profiles |
| Phase 3: Robustness | Probing operational boundaries and resilience | Deliberate variations in pH (±0.2), flow rate (±10%), temperature (±5°C) | Documented system suitability criteria and parameter tolerances |
| Phase 4: Validation | Formal statistical proof of analytical performance | Linearity, accuracy, precision (%RSD < 2%), LLOQ, matrix effects | Comprehensive validation report for audit readiness |
Phase 1: Scouting and Feasibility Screening
The scouting phase explores broad parameter spaces rather than chasing immediate analytical perfection. During this phase, analytical chemists evaluate fundamental chemical compatibility:
- Matrix Compatibility: Screening how the raw sample responds to thermal digestion, acid breakdown, organic precipitation, or liquid extraction.
- Separation and Retention Screening: For chromatographic methods, testing multiple stationary phase chemistries (e.g., C18, Phenyl-Hexyl, HILIC) across broad pH ranges (pH 2 to 10) to map retention patterns.
- Ionization and Detection Modes: For mass spectrometry and optical emission systems, evaluating ionization efficiency, isobaric interferences, and spectral emission lines.
Scouting establishes what is chemically feasible, allowing the laboratory to discard unworkable mechanisms early before committing time to fine-tuning.
Phase 2: Systematic Method Optimization
Once a workable analytical baseline is identified, optimization sharpens peak resolution, lowers baseline noise, and maximizes extraction recovery. Modern laboratories utilize Design of Experiments (DoE) to evaluate multiple interacting factors simultaneously rather than relying on one-factor-at-a-time guesswork.
During this stage, chemists optimize extraction yields and eliminate baseline drift. If an unexpected chromatographic anomaly occurs, applying targeted techniques for troubleshooting analytical method problems ensures that problems like peak fronting, tailing, or split peaks are solved at the root chemical level rather than masked by software integration tricks.
Phase 3: Robustness Testing and Operational Range Definition
A method that only works on one specific afternoon in the hands of a single expert scientist is not a valid method—it is a lab curiosity. Robustness testing deliberately introduces minor operational variations to identify the boundaries of method stability:
- Modifying mobile phase organic composition by ±2–5%
- Shifting buffer pH by ±0.2 units
- Varying column or spray chamber temperature by ±2–5°C
- Altering extraction hold times and centrifugation speeds
By testing these subtle shifts against standard system suitability benchmarks—such as maintaining a peak area relative standard deviation (%RSD) below 2%—the laboratory defines the Method Operable Design Region (MODR) in alignment with modern ICH Q14 guidelines.
Overcoming Technical Roadblocks in Difficult Matrices and Trace Quantitation
Developing methods for real-world samples means encountering complex chemical roadblocks. Pristine analytical standards in pure solvent rarely behave like dirty industrial extracts or biological fluids.
When standard methods hit a wall, evaluating the ultimate guide to outsourcing inorganic elemental analysis can help determine when external specialized sample preparation infrastructure is required.
How a Custom Method Development Laboratory Solves Complex Matrix Interferences
Matrix components can foul instruments, suppress detector ionization, and obscure trace target signals. Specialized laboratories deploy targeted sample preparation chemistries to solve these issues:
- High-Throughput Cleanup vs. Dilute-and-Shoot: In routine biological fluid or urine drug testing, basic dilute-and-shoot protocols frequently foul analytical columns in as few as 15 injections. By developing targeted solid phase extraction (SPE) or rapid enzyme/protein removal steps, specialized separation laboratories have demonstrated column life extensions exceeding 33x—reaching more than 500 injections while stabilizing backpressure.
- Multi-Analyte Extraction Consolidation: Analyzing complex analyte panels often requires separate workflows across different physical fractions. Tailored extraction methods can unify these paths. In specialized clinical testing, developing a single combined whole-blood extraction procedure reduced preparation time from over 3 hours down to roughly 45 minutes.
- Spectral and Isobaric Interferences: In trace elemental analysis, matrix elements can form polyatomic ions that share identical mass-to-charge ratios with target analytes (e.g., $^{40}\text{Ar}^{35}\text{Cl}^{+}$ interfering with $^{75}\text{As}^{+}$). Custom method developers use collision/reaction cells with specific cell gases (helium, hydrogen, oxygen) or alternative multi-element techniques to resolve these interferences cleanly.
Enhancing Sensitivity and Lower Limits of Quantitation (LLOQ)
Achieving low Lower Limits of Quantitation (LLOQ) requires an orchestrated effort across sample preparation and instrument physics. When target analytes exist at sub-parts-per-billion levels, custom development employs specialized strategies:
- Pre-Concentration Protocols: Using selective solid phase extraction cartridges or controlled solvent evaporation to concentrate trace analytes into micro-volumes without analyte loss.
- Chemical Derivatization: Attaching chemical tags to non-polar or poorly ionizing molecules to dramatically boost ionization efficiency and signal-to-noise ratios during spectrometry detection.
- Advanced Detection Selection: Matching the specific elemental or chemical profile to the optimal instrument platform. You can review a quick start guide to elemental analysis methods to see how techniques like ICP-MS, ICP-OES, and Particle Induced X-ray Emission (PIXE) differ in sensitivity and sample preservation.
Regulatory Validation Tiers and Defensible Method Transfer
Validation is not a monolithic check-the-box event. The level of analytical validation must match the intended end-use of the data, progressing along phase-appropriate tiers from early research to clinical or environmental submission.
Validation Tiers: Exploratory vs. RUO vs. GCP/GCLP
Understanding validation tiers prevents over-spending on early-stage discovery while ensuring full compliance when heading into regulatory filings:
- Exploratory Validation: Designed for early screening and proof-of-concept testing. Typically involves a single analytical run evaluating basic linearity, selective signal response, and single-level quality control (QC) precision.
- Research Use Only (RUO): Ideal for non-clinical candidate evaluation and internal decision-making. Requires testing across a minimum of three independent runs, evaluating three distinct QC levels (low, medium, high), baseline recovery, carryover, and short-term sample stability.
- GCP/GCLP/GMP Regulated Validation: Mandatory for clinical trials, release testing, and formal regulatory filings. Adhering to international frameworks such as ICH M10 for bioanalysis and ICH Q2(R2) for pharmaceutical quality, this tier assesses up to 18 distinct performance parameters. It demands comprehensive matrix factor testing across individual lots, dilution parallelism, long-term matrix stability, secondary instrument verification, and full QA audit trail reviews. For high-throughput clinical programs, leveraging standardized multi-site protocols helps ensure data consistency.
Selecting the Right Custom Method Development Laboratory for Trace and Bioanalytical Needs
Selecting an analytical partner is a major operational decision. Whether your program requires dedicated Full-Time Equivalent (FTE) staffing models for iterative research or transactional fee-for-service project scopes, specific partner capabilities are essential:
- Proven Regulatory Audit Track Record: Ensure the facility maintains verified quality management systems aligned with ISO/IEC 17025 or cGMP compliance. Reviewing accredited testing laboratory services that keep you audit ready highlights what documentation auditors inspect.
- Specialized Instrumentation Versatility: An effective partner should not attempt to force every problem into a single machine. They must offer diverse options—from chromatography and mass spectrometry to non-destructive elemental profiling. You can consult all about elemental analyzer options a comparison guide to evaluate instrument capabilities.
- Contemporaneous Documentation Culture: Method transfer often fails not because of chemical breakdowns, but because minor analyst habits—such as specific buffer dissolution orders, sonication times, or ambient hold conditions—went unrecorded in the lab notebook. A reliable development partner captures these subtle variables in real time.
- Specialized Matrices and Elemental Acumen: When screening for toxic contaminants or trace metals, evaluate their background through guides like how to choose the right heavy metal testing laboratory for your needs to ensure they possess the specialized digestion equipment needed to handle hazardous or recalcitrant materials. Academic and research programs exploring agricultural and plant metabolomics often refer to resources like the Center for Agricultural and Life Sciences Metabolomics (CALM) for specialized matrix extraction ideas.
Frequently Asked Questions About Custom Method Development
How long does custom method development typically take?
The timeline depends on matrix complexity, target analyte stability, and the required level of regulatory validation.
A straightforward feasibility and scouting phase on a well-behaved sample may take 2 to 4 weeks. However, developing a multi-analyte assay for complex matrices—incorporating optimization, robustness testing, stability profiles, and formal multi-run ICH validation—typically requires 2 to 4 months of iterative work, where long-term stability testing runs parallel to final protocol documentation.
What is the difference between method development, validation, and verification?
These three terms represent distinct milestones in an analytical method’s lifecycle:
- Method Development: The de novo process of establishing chemistry, sample extraction, separation conditions, and detection settings for an unvalidated or novel application.
- Method Validation: The experimental and statistical proof that a newly developed, non-compendial method meets all predefined performance requirements (specificity, linearity, accuracy, precision, limits of detection) for its intended regulatory purpose.
- Method Verification: The abbreviated process of demonstrating that an existing, officially validated compendial standard (such as a published USP or ASTM monograph) performs acceptably on your specific instrument and in your specific laboratory environment.
Why do analytical methods fail during inter-laboratory transfer?
Method transfer failures rarely stem from sudden changes in basic chemical laws; they fail because of unrecorded operational nuances.
Common culprits include differences in instrument dwell volume between instrument brands, subtle gradient delay differences, subtle differences in column packing lots, unrecorded laboratory temperature fluctuations, or incomplete reagent preparation instructions (such as adding acid to water versus water to acid). When a development laboratory maintains complete, contemporaneous documentation that captures failed trials alongside successes, receiving laboratories can reproduce the method without endless troubleshooting cycles. Technical teams frequently turn to resources like CUSTOM METHOD DEVELOPMENT to establish standard transfer protocols before shipping samples.
Conclusion
Analytical method development is the foundation that supports accurate product characterization, regulatory approvals, and manufacturing quality control. Forcing complex samples into ill-fitting compendial methods introduces operational blind spots, matrix interferences, and data integrity risks that can derail commercial timelines.
Partnering with an experienced development laboratory gives you fit-for-purpose assays designed specifically around your materials. At Elemental Analysis Inc. in Lexington, KY, we deliver over 40 years of specialized expertise in trace element identification, quantification, and speciation. As the world’s premier commercial Particle Induced X-ray Emission (PIXE) laboratory alongside comprehensive ICP-MS, ICP-OES, and chromatography capabilities, our team provides destructive and non-destructive testing with rapid turnarounds and competitive pricing.
Whether you need to quantify trace impurities in high-performance materials, solve complex matrix interferences, or develop audit-ready assays from the ground up, we are here to support your technical goals. Explore our full suite of analytical services or contact our team to discuss your project requirements with our specialized ICP-MS and elemental analysis services team today.